
The Prescription Drug User Fee Act (PDUFA) is a landmark piece of legislation that fundamentally reshaped the landscape of pharmaceutical development and regulation in the United States. Far more than a mere fee collection mechanism, PDUFA represents a significant instance of regulatory innovation, designed to accelerate the review process for new drugs and biologics by the Food and Drug Administration (FDA). Its establishment in 1992 marked a pivotal shift, introducing a system where pharmaceutical companies pay fees to supplement congressional appropriations, thereby funding the FDA’s drug review activities. This legislative framework, periodically reauthorized by Congress, has profound implications for how new technologies and innovations in medicine reach patients, influencing everything from research and development timelines to market access and public health outcomes.
The Genesis of a Regulatory Innovation
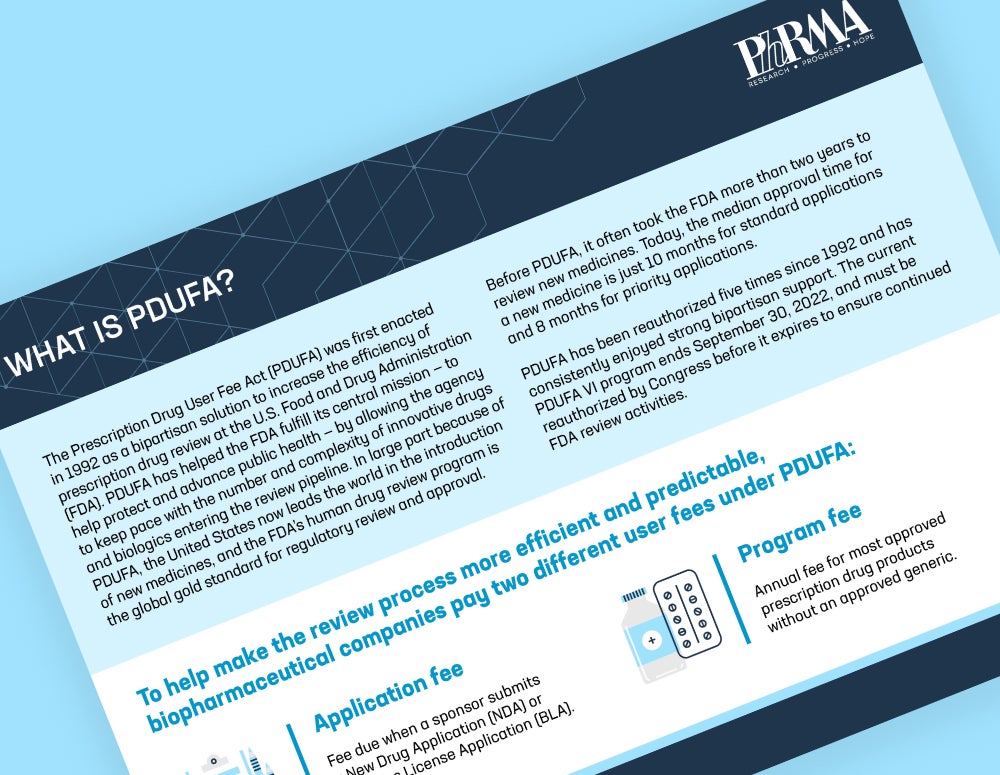
Before PDUFA, the FDA’s drug review process was notorious for its lengthy timelines, often spanning years. This “drug lag” was a growing concern for both pharmaceutical companies, which faced significant financial burdens from delayed market entry, and patient advocacy groups, who saw crucial treatments held up. The FDA, constrained by limited resources and an escalating number of complex drug applications, struggled to keep pace with scientific advancements and public demand.
Addressing the Drug Lag
The imperative behind PDUFA was clear: to enhance the efficiency and predictability of the drug approval process. The 1980s witnessed a growing frustration with the slow pace of drug availability, particularly during the AIDS epidemic, which underscored the urgent need for faster access to new therapies. This created a powerful coalition of industry and patient groups advocating for change. They argued that additional funding, directly tied to performance goals for drug reviews, could significantly reduce approval times without compromising safety and efficacy standards. PDUFA was conceived as a solution to this multifaceted problem, promising a more robust and responsive regulatory environment.
The User Fee Model Explained
At its core, PDUFA established a system where pharmaceutical and biotechnology companies applying for drug approval pay a “user fee” to the FDA. These fees are allocated specifically to funding additional FDA staff, technology upgrades, and other resources necessary to expedite the review of new drug applications (NDAs) and biologic license applications (BLAs). In return for these fees, the FDA commits to meeting specific performance goals for review times. These goals include target deadlines for reviewing applications, conducting facility inspections, and providing timely feedback to applicants. This quid pro quo arrangement created a unique partnership between the regulator and the regulated, fostering a shared interest in efficient and predictable outcomes. The funds generated by PDUFA significantly augment the FDA’s budget for drug review, enabling the agency to invest in necessary infrastructure, expertise, and scientific tools to evaluate cutting-edge pharmaceutical technologies.
PDUFA’s Impact on Pharmaceutical Technology & Innovation
PDUFA’s influence extends far beyond mere administrative efficiency; it has become an integral component of the ecosystem that fosters and brings pharmaceutical innovation to fruition. By creating a more predictable and often faster pathway to market, PDUFA inherently impacts strategic decisions in research and development, influencing the types of technologies pursued and the speed at which they are advanced.
Accelerating Drug Development
The primary, and most evident, impact of PDUFA has been the acceleration of the drug approval timeline. Reduced review times mean that innovative therapies can reach patients sooner, which is a powerful incentive for companies investing heavily in R&D. This predictability allows pharmaceutical innovators to better plan their clinical trials, manage financial investments, and forecast market entry. For highly specialized or breakthrough therapies, faster approval can be critical for patient outcomes and can also provide a competitive advantage, encouraging further investment in high-risk, high-reward technological avenues. The emphasis on defined review metrics has also pushed the FDA to adopt more efficient internal processes, leveraging data analytics and project management techniques to streamline evaluations of increasingly complex scientific dossiers.
Fostering Specific Technological Advances
PDUFA has, perhaps indirectly, helped to foster specific technological advancements within the pharmaceutical industry. For instance, the drive for efficiency encouraged by PDUFA means that companies are incentivized to submit well-organized, comprehensive data packages that leverage advanced data collection and analysis tools. Furthermore, the FDA’s enhanced resources, funded by PDUFA, have allowed it to invest in expertise required to evaluate novel technologies such as gene therapies, cell therapies, and personalized medicines. These advanced therapeutic modalities often present unique regulatory challenges, and PDUFA funds enable the FDA to recruit and train scientists with specialized knowledge in areas like genomics, bioinformatics, and advanced manufacturing processes. Without adequate regulatory capacity, the development and approval of these cutting-edge technologies would undoubtedly face greater hurdles.

Data, Efficiency, and Predictive Analytics
The iterative reauthorization of PDUFA (PDUFA I through VII and beyond) has seen an increasing focus on the use of data and technology to enhance regulatory efficiency. Subsequent iterations have incorporated provisions related to real-world evidence, patient-focused drug development, and advanced manufacturing technologies. This pushes both industry and the FDA towards more sophisticated data analytics, predictive modeling, and digital health tools to monitor drug safety and efficacy post-market. For example, the use of AI in drug discovery or advanced computational models for clinical trial design benefits from a regulatory environment that is increasingly adept at evaluating complex data sets and novel methodologies. PDUFA helps ensure the FDA has the computational infrastructure and skilled personnel to review submissions that increasingly rely on these advanced technological approaches.
Challenges and Evolving Regulatory Landscape
While PDUFA has largely been successful in its objectives, it also presents ongoing challenges and demands continuous adaptation to the rapidly evolving landscape of medical technology and public health needs.
Balancing Speed and Safety
One of the persistent challenges with PDUFA is the delicate balance between accelerating drug approvals and maintaining the highest standards of safety and efficacy. Critics occasionally raise concerns that the financial relationship between industry and the FDA could create a perception, if not a reality, of undue influence or pressure to approve drugs more quickly. However, the FDA has consistently maintained that its review decisions remain independent and scientifically rigorous, with PDUFA funds specifically earmarked for review-enhancing activities, not for approval outcomes. The reauthorization processes of PDUFA invariably involve extensive public hearings and stakeholder consultations to address these concerns and refine performance goals, ensuring that public health remains paramount.
Adapting to Emerging Technologies
The pharmaceutical and biotech sectors are characterized by relentless innovation. New therapeutic modalities, such as CRISPR-based gene editing, mRNA vaccines, and sophisticated AI-driven drug discovery platforms, present unprecedented scientific and ethical questions. PDUFA’s reauthorizations are crucial opportunities to ensure that the FDA’s capabilities keep pace with these emerging technologies. This often means allocating funds for research into new regulatory science, developing guidance for novel products, and fostering international collaboration on harmonized standards. The ability of the FDA to proactively engage with and develop regulatory pathways for these cutting-edge innovations is critical for their successful translation from lab to clinic. Each reauthorization must anticipate and address the technological shifts on the horizon, ensuring the regulatory framework remains agile and responsive.
PDUFA’s Broader Implications for Tech Regulation
PDUFA’s success in leveraging user fees to enhance regulatory capacity has prompted discussions about its potential as a model for other highly innovative and regulated industries.
A Model for Other Industries?
The PDUFA model, centered on industry funding for enhanced regulatory review capacity tied to performance metrics, has been observed with interest by other sectors grappling with complex technological innovation and regulatory bottlenecks. While direct application might be challenging due to differing industry structures and public interests, the core principle—that regulated industries can contribute to the funding of the regulatory agencies overseeing them, thereby improving efficiency and predictability—holds appeal. This could potentially apply to other areas of tech, where rapid innovation outpaces traditional regulatory frameworks, creating demands for specialized expertise and faster review times without compromising essential oversight.
The Future of Collaborative Regulation
PDUFA represents a unique form of collaborative regulation, where the industry provides resources, and the agency provides timely, high-quality review. This collaborative spirit is increasingly vital in a world of complex, rapidly evolving technologies. As technological advancements continue to accelerate across various sectors, the PDUFA model offers insights into how regulatory bodies can enhance their capabilities, attract specialized talent, and adapt to new challenges through structured engagement with the industries they oversee. It underscores the idea that effective regulation, particularly in high-tech fields, can be a shared responsibility that fosters innovation while safeguarding public interest. The evolution of PDUFA will continue to reflect the ongoing dialogue between technological progress, regulatory imperatives, and societal needs.